GMP & Pharmaceutical Manufacturing Expert
✓ Reviewed by: Pankaj Sharma - Quality Control Specialist
Reviewed for Quality Control accuracy, laboratory practices, analytical methods, and technical relevance
📅 Last Updated: November 13, 2026
Updated on November 2026: Corrected Taq names and cycling values, added PCR types, controls, troubleshooting, Ph. Eur. 2.6.7 QC uses, FAQ and references.
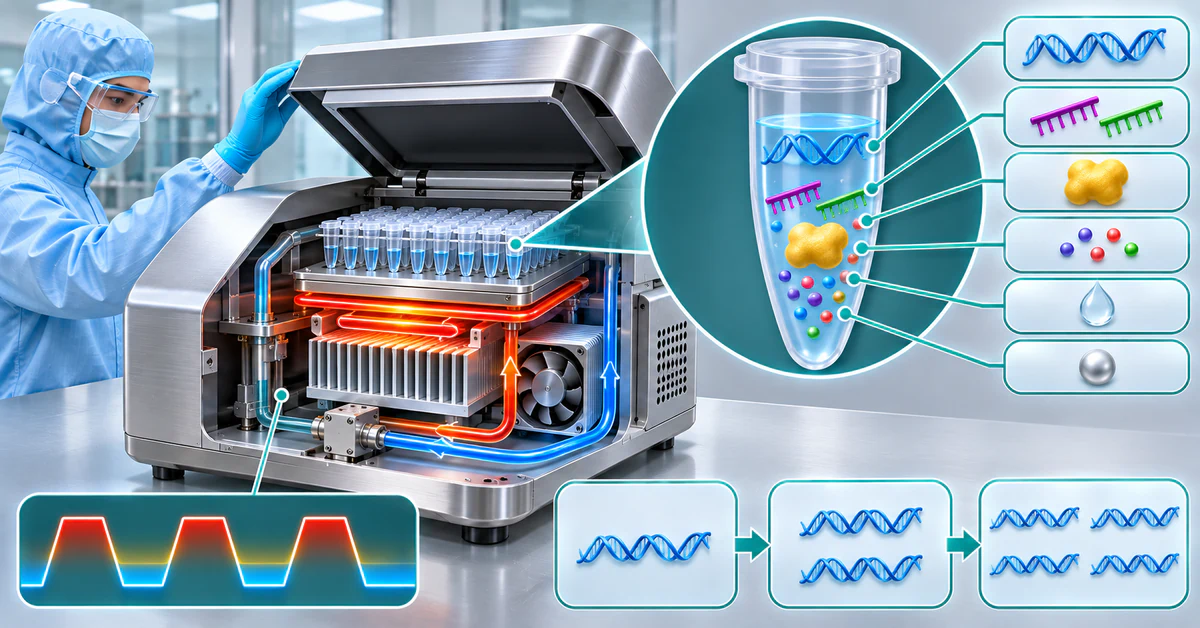
PCR is a laboratory technique that makes millions to billions of copies of a chosen DNA segment. Primers mark the segment, and a thermal cycler repeats denaturation (about 94 °C), annealing (about 45–60 °C) and extension (72 °C) for 30 to 40 cycles using Taq polymerase.
What is polymerase chain reaction (PCR)?
Polymerase chain reaction (PCR) is a technique used to make many copies of a small fragment of DNA. It emerged in the mid-1980s. Kary Mullis conceived the method in 1983 and received the 1993 Nobel Prize in Chemistry for it. RNA can also be studied, but it must first be converted to complementary DNA (cDNA) before PCR, as described under reverse transcription PCR (RT-PCR) below.
The name has two parts: a polymerase is an enzyme that builds a new DNA chain, and a chain reaction is a reaction in which each round of copying provides the template for the next round.

In each cycle the quantity of DNA doubles. One molecule therefore becomes 2, 4, 8, 16, 32, 64, 128 and so on (2^n). About 30 ideal cycles give over 1 billion copies from one starting molecule, and a typical run has 30 to 40 cycles. These copies are used to diagnose viral infections, in forensic investigation and in genetic research, and also in pharmaceutical quality control (see the applications section below).
Components required for PCR
A PCR reaction mixture needs the following components:
- The target DNA (template), which can be genomic DNA, cDNA or a plasmid.
- Two primers, each about 20 nucleotides long, complementary to the sequences next to the DNA segment of interest.
- A thermostable DNA polymerase, usually Taq polymerase.
- Free deoxynucleotides (dNTPs), which are the building blocks of the new strands.
- Reaction buffer containing Mg2+, which acts as a cofactor for the polymerase.
- A thermal cycler.
Taq polymerase
Taq polymerase comes from Thermus aquaticus, a bacterium that lives in hot springs. The enzyme is heat stable (half-life about 40 minutes at 95 °C), so it survives the high temperature of each cycle. It typically has its optimum activity at about 75–80 °C, where it adds about 150 nucleotides per second. In practical PCR, about 1 kb per 30–60 seconds is typical, and these values vary by manufacturer. Taq has no proofreading activity, and high-fidelity enzymes are therefore used when copies are needed for cloning.
Primers
DNA polymerase cannot start a new strand and can only extend an existing primer, so each PCR needs two primers, and the primers also select the part of the DNA that is amplified. Good primers meet these design rules:
A length of 18–25 nucleotides.
A Gas Chromatography (GC) content of 40–60%.
Matched melting temperatures (Tm).
No tendency to form dimers.
The primer concentration in the reaction is 0.05–1.0 µM.
Thermal cycler
The thermal cycler heats and cools the tubes automatically, so the reaction passes through the same temperature steps in every cycle.
| Component | Role | Final concentration |
|---|---|---|
| Template DNA | Source of the target sequence | Set by the assay |
| Primers (forward and reverse) | Mark the start of each new strand | 0.05–1.0 µM |
| Taq polymerase | Synthesises the new strands | Set by the supplier |
| dNTPs | Building blocks of DNA | Set by the supplier |
| Buffer with Mg2+ | Maintains conditions, Mg2+ is the cofactor | Optimised for each assay |
Principle and steps of polymerase chain reaction (PCR)
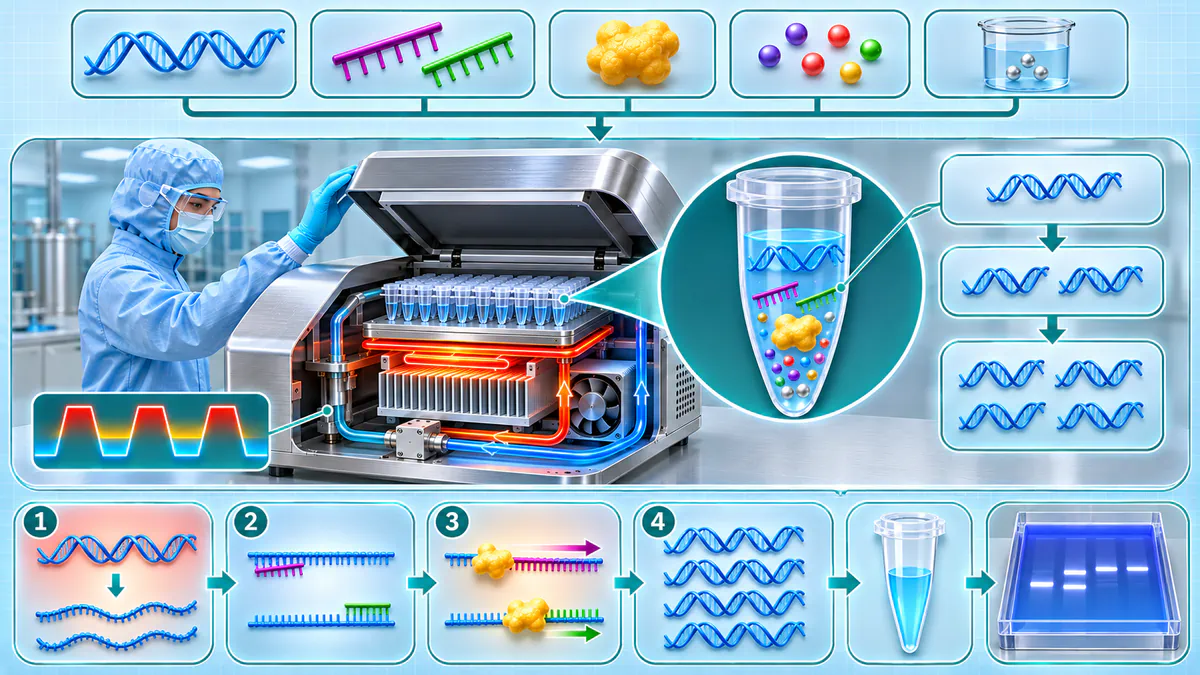
High temperature separates the two DNA strands, and a lower temperature lets the primers bind to them. The polymerase then extends each primer, so both strands are copied and the quantity of DNA doubles in every cycle. An initial denaturation at about 94 °C is carried out once before cycling starts. The steps below are then repeated for 30 to 40 cycles in an automated cycler.
1. Denaturation at about 94 °C for 15–60 seconds

The high temperature (94–98 °C) breaks the hydrogen bonds between the bases, and the double-stranded DNA melts open into two single strands. The strands are separated without being cut.
2. Annealing at about 45–60 °C (set from primer Tm)

The mixture is cooled so that the primers can bind to the single strands. The annealing temperature is set from the primers used and is 3–5 °C below the primer Tm. The primer and the template continually form and break hydrogen bonds, and primers that fit perfectly stay bound longest. The polymerase attaches to this short double-stranded region and starts copying, and after a few bases are added the bond becomes strong enough to stay intact.
3. Extension at 72 °C
At 72 °C (70–75 °C) the polymerase works at its best. It reads the template from 3′ to 5′ and adds complementary dNTPs to the 3′ end of the primer, so the new strand grows from 5′ to 3′. Primers with no exact match loosen at this temperature and do not extend. The extension time is set by the length of the product (amplicon), at about 1 minute per kb for Taq.
| Step | Typical setting | Example programme |
|---|---|---|
| Initial denaturation | About 94 °C, once | Before cycle 1 |
| Denaturation | About 94 °C, 15–60 s | 94 °C, 30 s |
| Annealing | About 45–60 °C | 54 °C, 45 s |
| Extension | 72 °C, about 1 min per kb | 72 °C, 2 min (for a 2 kb product) |
| Cycles | 30–40 | 30 |
Exponential amplification
Both strands are copied in every cycle, so one starting copy gives 2 copies after one cycle, 4 after two cycles and 8 after three cycles. After x cycles the copies are given by 2^x (2 to the power x). This is the ideal maximum and not the real yield. The real yield is lower and reaches a plateau as the reagents are used up.
| Cycles | Ideal copies from one molecule |
|---|---|
| 10 | 1,024 |
| 20 | 1,048,576 |
| 30 | Over 1 billion |
PCR full form is Polymerase Chain Reaction
Polymerase chain reaction is a process that is used to create several copies of a small fragment of DNA. PCR is used in the identification of types of infections, crime investigation, molecular biology, and genomics research
Its basic Principle is based on the replication of particular DNA sequences in vitro using DNA polymerase.
Thermos Aquaticus enzymes
READ ALSO
Voltammetry and Polarography Explained
Voltammetry is a time-dependent potential applied to an electrochemical cell.
Types of PCR
The basic method has been modified for different purposes, and the main types are compared below.
| Type | Key feature |
|---|---|
| Conventional (end-point) PCR | The product is read at the end of the run, for example on a gel |
| RT-PCR | RNA is first converted to cDNA, either in one tube (one-step) or in separate reactions (two-step) |
| qPCR (real-time PCR) | Fluorescent dyes or probes are measured in every cycle, and the Cq (Ct) value and a standard curve give the starting amount |
| Multiplex PCR | Several targets are amplified in one tube with different primer pairs |
| Nested PCR | A second round with inner primers amplifies the product of the first round |
| Hot-start PCR | A modified Taq stays inactive until heated, which reduces primer dimers and non-specific products |
| Long-range PCR | Long amplicons are produced with suitable enzymes |
| Digital (droplet) PCR | The reaction is partitioned, and the target is counted directly |
Conventional PCR therefore only shows the product at the end, while qPCR measures fluorescence in each cycle to quantify the starting amount. A melt curve can also be run after qPCR to check the product.
Critical parameters of a PCR run
The parameters below are typical ranges, and they can vary between manufacturers and assays.
| Parameter | Typical range | Why it matters | How it is monitored |
|---|---|---|---|
| Denaturation | About 94 °C (94–98 °C), 15–60 s | Strands must separate fully | Programmed cycler profile |
| Annealing | 45–60 °C, 3–5 °C below Tm | Controls primer specificity | Programmed profile, gel or melt curve |
| Extension | 72 °C (70–75 °C), about 1 min per kb | Allows complete products | Programmed profile, product size on gel |
| Cycles | 30–40 | Sets the yield | Programmed profile |
| Primer concentration | 0.05–1.0 µM | Affects yield and primer dimers | Preparation record |
| Mg2+ | Optimised for each assay | Acts as the polymerase cofactor | Preparation record, test run |
PCR controls and contamination prevention
Every run should include the following controls:
- A no-template control, which shows contamination.
- A positive control, which shows that the reaction works.
- An internal (inhibition) control, which shows whether the sample inhibits amplification.
Contamination is prevented by the following measures:
- Separate pre-PCR and post-PCR areas with a one-way workflow, so that amplified product never returns to the set-up area.
- Dedicated pipettes.
- Filter or positive-displacement tips.
- Gloves.
Troubleshooting PCR
This section lists the common faults of a PCR run, their likely causes and the fixes.
| Problem | Likely cause | Fix |
|---|---|---|
| No product | Poor primer design or Tm, unsuitable Mg2+, annealing temperature or extension time | Check the primer design and Tm, then optimise Mg2+, annealing temperature and extension time, keeping primers at 0.05–1.0 µM |
| Non-specific bands or primer dimers | Primers binding at the wrong sites or to each other | Use hot-start Taq, set up on ice and raise the annealing temperature in 2 °C steps |
| Contamination | Carry-over of product or template | Use a dedicated set-up area, dedicated pipettes and filter tips |
Applications of PCR, including pharmaceutical QC
PCR is used in the diagnosis of viral infections, in forensic investigation and in genetic research. In pharmaceutical quality control, it is used for the following purposes:
- Mycoplasma testing.
- Measurement of residual host-cell DNA.
- Detection of adventitious viruses.
- Microbial identification.
- Cell-line authentication.
Like any analytical test, such as the limit test for sulphate, a PCR method must be validated before it is used for release decisions.
Validated nucleic acid amplification techniques such as PCR are accepted as an alternative to culture and indicator-cell methods for mycoplasma testing under revised Ph. Eur. 2.6.7 (version 12.2). The method must meet these requirements:
- A limit of detection (LOD) of 10 CFU/mL or fewer than 100 genomic copies/mL.
- Validation in the user’s own product matrix.
- Internal, positive (near the LOD) and negative controls in every run.
The revision was first published for public comment in 2022, and it is reported to be mandatory from April 2026.
The International Council for Harmonisation (ICH) guideline ICH Q5A(R2), reported as adopted on 1 November 2023, covers molecular assays such as PCR in the viral safety evaluation of biotechnology products.
Digital PCR (ddPCR) gives absolute quantification without a standard curve, shows lower variability than qPCR and tolerates inhibitors better. It is therefore used to measure residual host-cell DNA in biologics and gene therapies.
<!– Accuracy flag (Applications): the Ph. Eur. effective date and LOD come from a vendor summary, the ICH Q5A(R2) date from a search result, and neither is confirmed on the EDQM or ICH site. The Food and Drug Administration (FDA) limit for residual host-cell DNA per dose is not stated because it is not confirmed on fda.gov. –>
Advantages, limitations and automation
The main advantages of PCR are speed, high sensitivity and the ability to work with tiny samples. Its limitations are as follows:
- The sequence of the target must be known to design primers.
- Contamination can cause false positives.
- Taq polymerase has no proofreading, so errors can enter the copies.
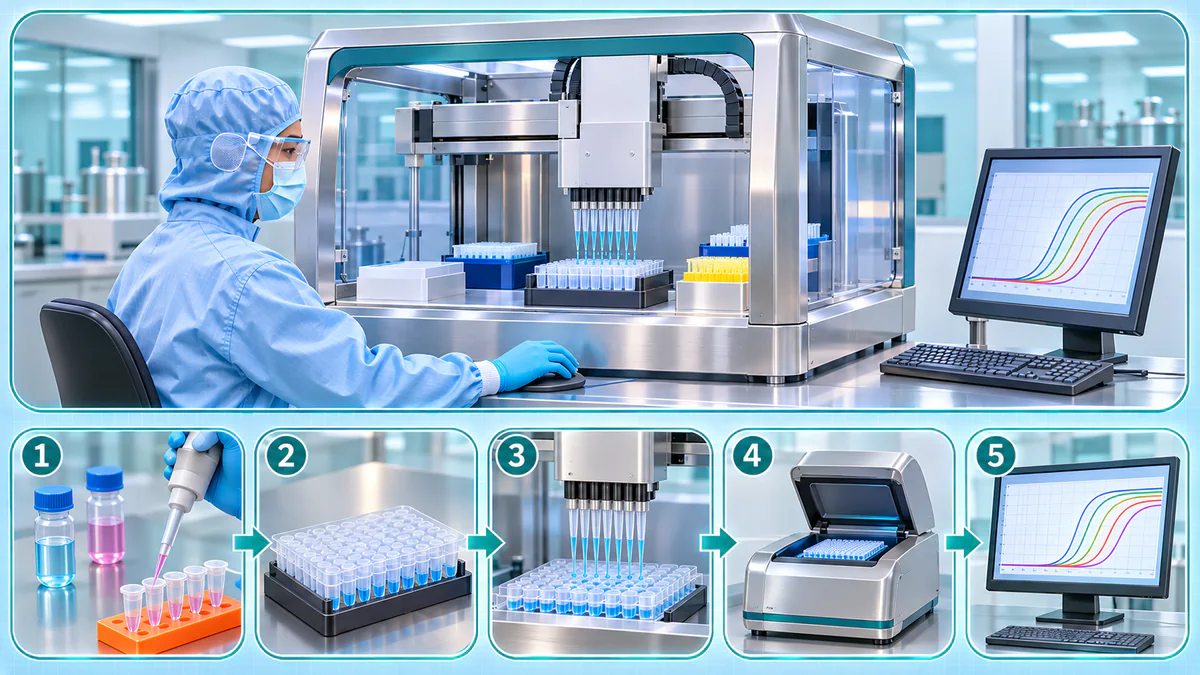
Automated and point-of-care systems are now in use. The AMDI Fast PCR instrument with a 4-plex assay for RSV, influenza A/B and SARS-CoV-2 gives results in under 10 minutes and holds FDA 510(k) clearance. Another system combines extraction and RT-PCR with robotic liquid handling.
In regulated laboratories, automated qPCR systems must be validated, and accurate liquid handling is needed because pipetting errors cause inconsistent Cq values. The electronic records of such systems require controls such as audit trails and electronic signatures (21 CFR Part 11).
Frequently Asked Questions
Who invented PCR?
Kary Mullis conceived PCR in 1983, and the technique came into general use in the mid-1980s. He received the 1993 Nobel Prize in Chemistry for the invention. The method spread quickly once a heat-stable polymerase removed the need to add fresh enzyme after every heating step.
Why is a thermostable polymerase needed?
The reaction is heated to about 94 °C in every cycle, which would destroy an ordinary polymerase. Taq polymerase from Thermus aquaticus survives this temperature, so it can be added once at the start and still work through all 30 to 40 cycles.
What is hot-start Taq?
Hot-start Taq is a modified Taq polymerase that stays inactive until the mixture is heated. Because it cannot extend primers during set-up at lower temperatures, it reduces primer dimers and non-specific products. Setting up the reaction on ice has a similar purpose.
Can PCR replace culture for mycoplasma testing?
Under revised Ph. Eur. 2.6.7, a validated nucleic acid amplification method can be used instead of culture and indicator-cell methods. The method must reach the required limit of detection, must be validated in the product matrix and must run with internal, positive and negative controls.
What is digital PCR used for?
Digital PCR is used for absolute quantification of rare targets, such as residual host-cell DNA in biologics. The reaction is divided into many partitions, and the target is counted directly, so no standard curve is needed and the result is less affected by inhibitors.
References
- Polymerase Chain Reaction, NHGRI Genetics Glossary
- Polymerase Chain Reaction guide, SnapGene
- Taq PCR Kit Troubleshooting Guide, New England Biolabs
- Compliant Mycoplasma Testing: New Regulatory Standard for NAT Methods, Sartorius
- Revision of Chapter 2.6.7 of the European Pharmacopoeia published for Comment, GMP-Compliance.org
- Quantitation of Residual Host Cell DNA in rAAV Using ddPCR, PMC
- Point-of-Care PCR Panel Detects RSV, Influenza, and SARS-CoV-2 in Minutes, LabMedica
- Fully Automated Detection System Revolutionizes Point-of-Care Testing, LabMedica
- How Automation is Transforming PCR Accuracy, Efficiency, and Scalability, BRAND
Naresh Bhakar is the Founder and Author at Pharmaguddu.com, bringing his extensive expertise in the field of pharmaceuticals to readers worldwide. He has experience in Pharma manufacturing and has worked with top Pharmaceuticals. He has rich knowledge and provides valuable insights and data through his articles and content on Pharmaguddu.com. For further inquiries or collaborations, please don’t hesitate to reach out via email at [email protected].