Quality Control Specialist | Industry Experience
✓ Reviewed by: Pankaj Sharma - Quality Control Specialist
Reviewed for Quality Control accuracy, laboratory practices, analytical methods, and technical relevance
📅 Last Updated: November 7, 2026
The potential areas of deviations in pharmaceutical manufacturing are materials, equipment, process, practices, documentation practices, test methods and specifications. A deviation is any departure from an approved procedure, parameter or specification. It must be documented and investigated, and its impact on product quality and compliance must be reviewed.
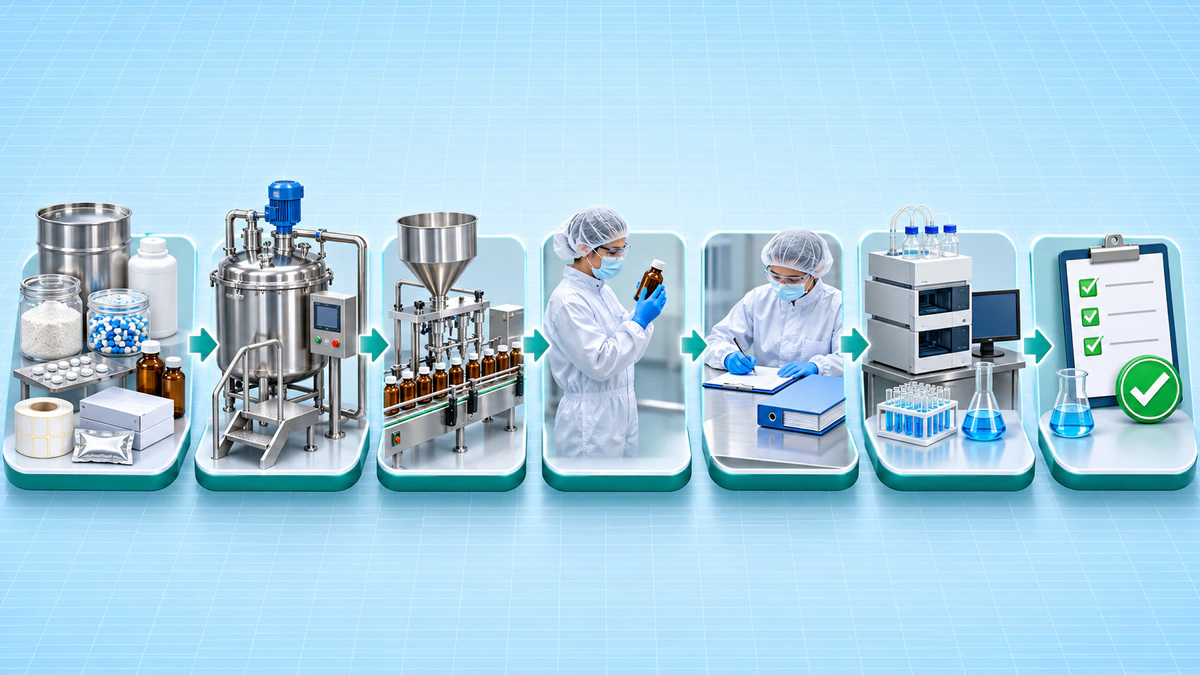
The list of potential areas of deviations in pharmaceutical manufacturing covers materials, machines, processes, practices, documentation, test methods and specifications. A deviation is any departure from an approved procedure, process parameter, specification or instruction that can affect product quality or GMP compliance. It must be documented and investigated. The types of deviations in pharmaceuticals are described in a separate post.
List of Potential Areas of Deviations in Pharmaceutical
The potential areas of deviations in pharmaceutical manufacturing are given below. Each area is described in the sections that follow.
- Materials
- Equipment
- Process
- Practices
- Documentation Practices
- Test Methods
- Specifications
Materials
The following must meet their quality specifications before use in manufacturing:
- Raw materials
- Components
- Labels
- Packaging materials
Sometimes an unreleased material reaches manufacturing. Sometimes an unapproved labeling component is used in the labeling or packaging operation.
In either case, Quality Assurance (QA) must review the impact on the quality and compliance status of the product in detail before deciding the disposition of the batch.
Equipment
The sudden breakdown or malfunction of equipment is an unplanned deviation that can have an adverse impact on quality.
Equipment used in manufacturing, packaging or testing after its calibration due date compromises the compliance status of the product. A calibration deviation may show no apparent effect on the quality, safety or efficacy of the drug substance, but it still has compliance implications.
Replacing equipment with similar or comparable equipment in the middle of a manufacturing run is another possible deviation. The production team must document and justify the change when it happens, and the QA team must approve it beforehand.
Process
Deviation from process parameters and from process conditions can affect the product.
The following must stay within preset limits:
- Machine setup
- Mixing speed
- Drying temperature
Any departure from these limits is a deviation. It must be recorded, and the depth of the investigation depends on the impact on product quality, not on the distance from the validation data alone.
Production staff must maintain process conditions such as:
- Temperature
- Humidity
- Microbial quality of the environment
Any departure from their set limits is a deviation and could affect product quality. Normal variation within the limits is not a deviation.
Practices
Written procedures must be followed strictly, and a failure to do so is a deviation. Under 21 CFR 211.100, production and process-control procedures must be written and followed, and any departure must be recorded and justified. Schedule M and EU GMP set the same expectation.
For example, if staff do not follow the sampling procedure, the sample becomes suspect. Test data from that sample cannot support any quality decision.
Such a deviation requires re-sampling and retesting of materials, along with other corrective actions, including retraining of the employees. The audit check list for quality assurance can be used to review these practices.
Documentation Practices
Schedule M requires quality-related documents to be prepared, reviewed, approved and controlled under written procedures. The same expectation appears in EU GMP Chapter 4 and WHO GMP. The list of quality assurance SOPs shows the procedures that govern such documents.
These documents must be used in a consistent manner. If they are not, the inconsistency is a deviation that has to be explained.
Poor documentation practice can occur in any department, so every employee must be trained and retrained on it.
Test Methods
Analytical test methods are qualified for a given drug substance or product and a set of instruments. Under International Council for Harmonisation (ICH) Q2(R2), an analytical procedure is validated to show it is suitable for its intended use, so any departure from the approved procedure needs a documented investigation and justification before the result is used.
Any deviation from the approved analytical procedure needs a detailed investigation and also a sound scientific justification, and QA accepts the data for a quality decision only after both are done.
Some deviations are minor, yet these should be documented. Others can lead to a re-evaluation of the analytical method or the instrument used.
Sometimes a test must deviate from the approved method because the compendium has changed. In that case, the organization has not yet reviewed and approved the updated written procedure.
Specifications
Finished product specifications are based on sound scientific data and are a direct result of the behavior of the drug substance or product during development.
Raw material specifications follow the current pharmacopoeia, such as Indian Pharmacopoeia (IP), British Pharmacopoeia (BP) or United States Pharmacopeia (USP), or the approved supplier specification. Packaging component specifications follow the same basis.
A deviation from an approved specification means the material fails. QA then rejects the material or sends it for rework.
Frequently Asked Questions
The answers below summarise the points on deviations in materials, equipment, practices, test methods and specifications described above.
The areas are materials, equipment, process, practices, documentation practices, test methods and specifications. A departure from an approved procedure, parameter or specification in any of these areas is a deviation. It must be documented and investigated, and its effect on product quality and compliance must be reviewed.
Use of equipment in manufacturing, packaging or testing after its calibration due date compromises the compliance status of the product, even though the effect on the quality, safety or efficacy of the drug substance may not be apparent. Compliance implications remain. It must therefore be handled as a deviation.
Yes. A mid-run replacement of equipment with a similar or comparable unit is a possible deviation. A planned replacement is a change, not a deviation: it needs QA approval through change control before it is carried out, and its impact is evaluated afterwards. An unplanned swap, for example after a sudden breakdown, is a deviation. The production team must document and justify it at the time it happens, and QA investigates and assesses it, with approval afterwards and not beforehand.
Written procedures must be followed strictly, so any departure from the sampling procedure is a deviation. The sample then becomes suspect. Test data from it cannot support any quality decision, and re-sampling, retesting and other corrective actions are then required, including retraining of employees.
A detailed investigation and a sound scientific justification are needed for any deviation from the approved analytical procedure. Only after both are complete does QA accept the data for a quality decision. Minor deviations should still be documented, and others can lead to a re-evaluation of the method or the instrument.
A deviation from an approved specification means the material fails. QA then rejects the material or sends it for rework. Before use in manufacturing, raw materials, components, labels and packaging materials must meet their quality specifications.
References
- 21 CFR 211.100, US Code of Federal Regulations (eCFR)
- Schedule M, CDSCO, Drugs Rules, 1945
- EU GMP, European Commission, EudraLex Volume 4
- WHO GMP, World Health Organization
- Indian Pharmacopoeia, Indian Pharmacopoeia Commission
- British Pharmacopoeia, British Pharmacopoeia Commission
- USP, United States Pharmacopeia (USP–NF)
Pankaj Sharma — Quality Control Specialist
Pankaj Sharma is a pharmaceutical QC professional who reviews our technical content for GMP accuracy, laboratory practices, and regulatory relevance.