Quality Control, Quality Assurance Specialist | Industry Experience
✓ Reviewed by: Pankaj Sharma - Quality Control Specialist
Reviewed for QC, QA accuracy, laboratory practices, analytical methods, and technical Documents
📅 Last Updated: November 14, 2026
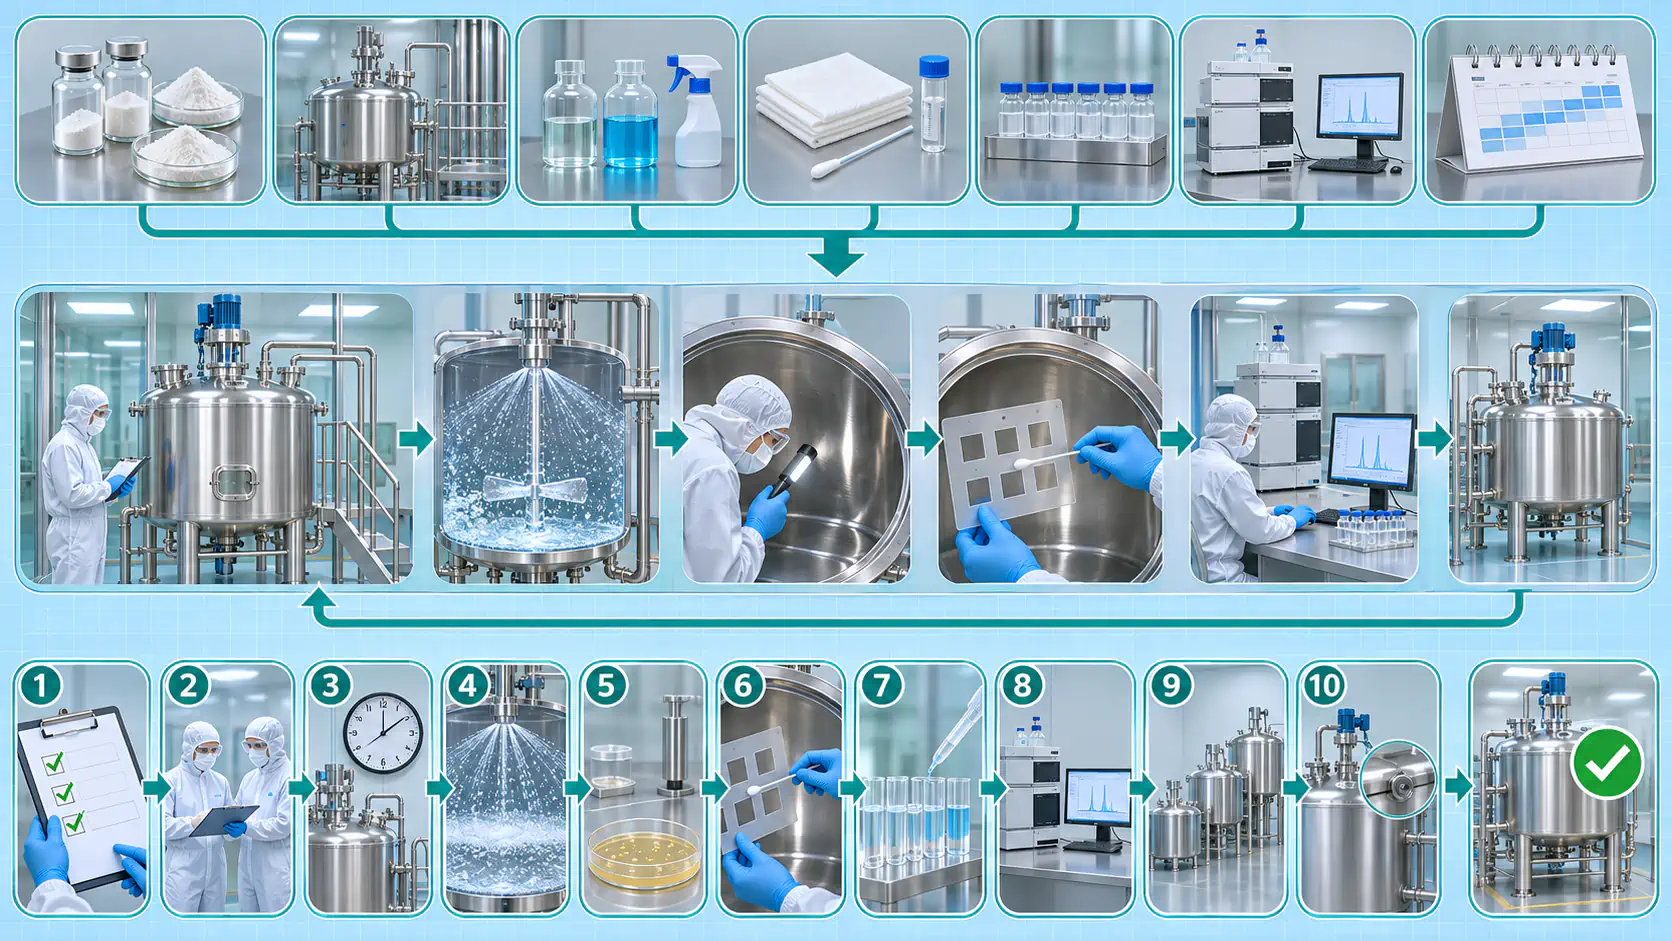
A cleaning validation protocol is a pre-approved plan that describes how a cleaning procedure will be tested and what results will count as acceptable. The Pharmaceutical Inspection Co-operation Scheme (PIC/S) document PI 006-3 defined cleaning validation as documented evidence that an approved cleaning procedure produces equipment suitable for processing pharmaceutical products. The aim is a reliable procedure. Routine analytical monitoring can then be omitted or kept to a minimum.
What principle does a cleaning validation study follow?
The principle is simple. Equipment is cleaned by a fixed written procedure, and measured residues of the previous product, the cleaning agent and microorganisms are then compared with limits that were fixed before the study. If every result meets the limits, repeatedly, the procedure is considered validated.
Three ideas support this principle. First, limits for carryover must rest on a toxicological evaluation. This is documented in a risk assessment (Annex 15, clause 10.6). Second, the study covers the worst case, such as the least soluble or most potent product and the hardest-to-clean equipment. Third, the sampling and analytical methods must be able to find the residue, so recovery from each surface material has to be shown.
The FDA inspection guide of 1993 adds that a negative result only means that the residue is below the detection limit of the method. It does not prove that no residue exists.
What does a protocol contain, and in what order is the study done?
| Title of Contents |
| Objective |
| Scope |
| Responsibility |
| Procedure |
| Sampling Plan |
| Sampling Points |
| Test/Method |
| Acceptance Criteria |
| ANNEXURE A – Cleaning Validation Test Request Form |
| ANNEXURE B – Cleaning Validation Test Report |
| ANNEXURE C – Cleaning Validation Test Report |
1.0 OBJECTIVE: To assure that the cleaning procedure for Equipment removes residues to the extent of compliance with the pre-determined acceptance level. (Sifter As an Example)
2.0 SCOPE: This protocol applies to the Pharmaceutical Manufacturing Facility.
3.0 RESPONSIBILITY: It is a joint responsibility of the Quality Control, Production, and Quality Assurance departments. The detailed responsibility matrix is given below
- Protocol Preparation- Executive QA
- Protocol Approval- Head QA
- Protocol Execution – Head Production
- Sampling as per Protocol – Executive QA
- Testing as per Protocol- Executive QA
- Test result view And approval- Manager QA
- Validation Data Compilation- Executive QA
- Validation final approval- Head QA
4.0 PROCEDURE
Clean the equipment as per SOP “Cleaning of Equipment”.
Sample from the specified sampling points (6.0) as per the sampling plan (5.0).
5.0 SAMPLING PLAN
- Sample from first rinse (after washing with raw water).
- Sample from second rinse (after cleaning with 0.1% liquid soap solution and raw water).Sample from final rinse of purified water.
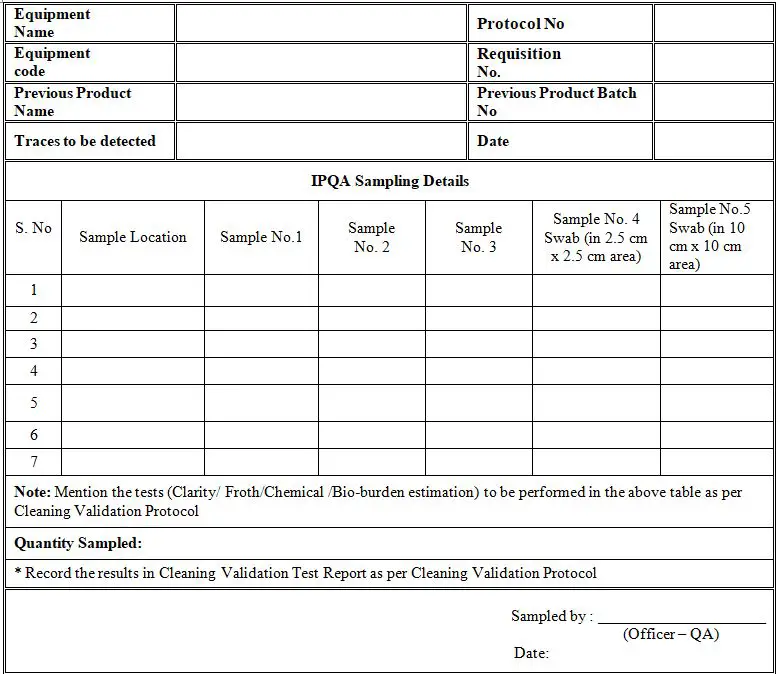
- Swab (in 2.5 cm x 2.5 cm area) for chemical analysis.
- Swab (in 10 cm x 10 cm area) for bio-burden.
6.0 SAMPLING POINTS
- Upper Hopper
- Sieve used for the batch
- Powder discharge assembly
Note: Sampling points may differ from equipment to Equipment (We used Shifter for Reference)
7.0 TEST / METHOD
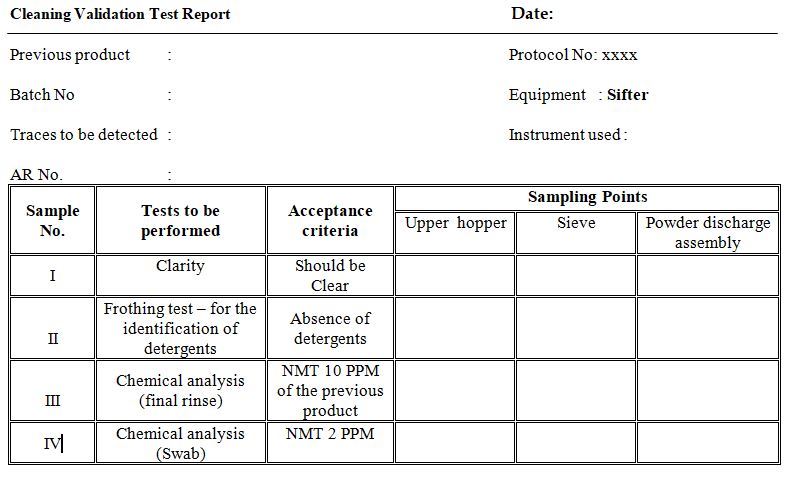
- Inspect visually for the absence of previous products in the equipment (sample I)
- Determine the absence of detergents by the froth method (sample-II ).
- Determine the concentration of active ingredient of the previous product in the final rinse (sample III).
- Determine the concentration of active ingredient of the previous product in the swab (sample IV).
- Report and record the above test (1,2,3 and 4) results in ANNEXURE – B
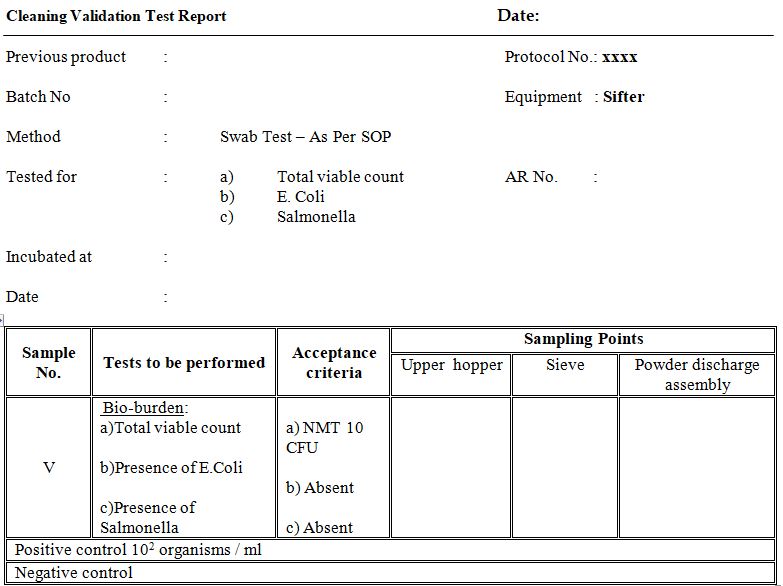
- Determine the bio-burden(sample V) and record the results in ANNEXURE– C
8.0 ACCEPTANCE CRITERIA
- No traces of the powder shall be visible on any part of the machine (sample-I)
- Detergent should not be identifiable by frothing test. (Sample-II).
- The concentration of the previous product should not be more than 10 PPM in the final rinse i.e. (sample III)
- The concentration of the previous product should not be more than 2 PPM in the swab (i.e. sample IV)
- Bio-burden limits are as below (sample V)
- Total CFU – NMT 10 CFU
- E. Coli -absent
- Salmonella – absent
What problems occur, and how are they fixed?
Out-of-limit residue result.
A result above the limit is a deviation. The QA officer starts an investigation, and the cause is corrected in the cleaning procedure, which is then re-studied. Repeated cleaning and retesting until the result passes is not acceptable under Annex 15, Annex 15, and the FDA guide says constant retesting shows that the process is not validated. The acceptance criteria must also not be changed after the results are seen.
Low or variable swab recovery.
The swab technique, the solvent or the surface material is reviewed, and the recovery study is repeated before the cleaning study continues. A trained sampler using a template reduces the variation.
High results only for hard-to-reach areas.
The sampling locations are reviewed, because rinse samples can reach inaccessible parts but depend on solubility, and a swab and rinse combination can then be used.
Residue after a long dirty hold time.
The hold time is shortened, or the cleaning procedure is strengthened and validated again at the longest hold.
Microbial growth after storage.
Equipment must be dried before storage and no stagnant water may remain, and the clean hold time is then reviewed.
Detergent detected in the final rinse.
The rinsing steps are reviewed. The FDA guide states that detergents should be easily removable, otherwise a different detergent should be selected.
Annexure- A (Cleaning Validation Test Request Form)
Annexure- B (Cleaning Validation Test Report)
Content of ___________________________in PPM
Area of sample x weight taken x dilution factor x 1000
Area of std. 100
Note: Attach calculation sheet and chromatogram.
Conclusion / Recommendations:
_______________________________________________________________________________________
Tested by: ______________ Approved by: ________________
(Analyst – QC) (Manager – QC)
Date:: Date:
Annexure – C (Cleaning Validation Test Report)
Analyzed by: _______________ Approved by: ________________
(Microbiologist) (Manager – QC)
Date: Date:
Frequently Asked Questions
Product-contact surfaces are the main focus. Non-contact parts to which product can migrate, such as seals, flanges, mixing shafts, oven fans and heating elements, should also be considered, as PI 006-3 noted.
The FDA guide calls direct surface sampling the most desirable method and accepts rinse sampling as the second type. A rinse covers a larger area and reaches parts that cannot be dismantled. Using both together gives the fullest picture.
A limit based on a toxicological evaluation, normally the HBEL with a MACO calculation, replaces fixed ppm values. Annex 15 asks for the justification to be documented with supporting references in a risk assessment.
Revalidation is triggered by a change in the cleaning procedure, equipment, product range, cleaning agent or batch size, as assessed through change control. The protocol states these triggers in advance.
Yes. The detergent composition should be known, and the supplier should notify any formulation change, because a changed detergent can alter residue testing and removal.
Pankaj Sharma — Quality Control Specialist
Pankaj Sharma is a pharmaceutical QC professional who reviews our technical content for GMP accuracy, laboratory practices, and regulatory relevance.